In pharma content localization, a global campaign rarely arrives in the US ready to use. One approved HCP email pack may need changes to claims language, product information, risk statements, references, brand terminology and channel format. Some global sections cannot travel as written.
Your global campaign may be ready while the US materials are still moving through adaptation and review. Launch dates slip. People rebuild sections that already exist elsewhere. Teams may lack a simple view of which version contains which change.
Why pharma content localization slows down in the US
The global source passes through several hands before it can be used in the US. The local commercial team adapts it. Medical, legal and regulatory specialists check the claims, supporting evidence, risk presentation and consistency with FDA-required labeling. Brand owners review consistency. An agency may sit between them.
The work becomes harder when the source or label changes. A revised safety statement may affect emails, banner ads, detail aids and other approved assets. Teams first need to find every affected version and check whether the old wording remains.
The workload is growing. Veeva reported that US content production rose 29% in 2023 compared with the previous year. Before formal review even starts, preparation can add anywhere from five to 150 days to each asset.
Put a number on the current process
There is no reliable industry figure for the full cost of adapting global content for the US. Each company has a different asset volume, agency model and review process. A simple internal calculation gives a more useful answer.
40 assets per quarter × 3 preparation hours = 120 preparation hours per quarter. Saving one hour on each asset would free up 40 hours per quarter. At an illustrative blended cost of $100 per hour, that equals $4,000 per quarter, or $16,000 per year if the same volume recurs in all four quarters.
The calculation excludes agency fees and the value of launching earlier. This is a planning example, not a promised saving. Replace each input with real operational data. The result becomes a clear baseline for deciding whether automation is worth testing.
The value isn’t limited to labor cost. Commercial teams get more time for campaign planning and performance analysis. Reviewers can focus on claims, evidence and risk presentation instead of preventable terminology or formatting errors.
A concrete global-to-US example
Consider a global team that has approved a two-email HCP sequence for a new oncology therapy. The US version needs claims that are consistent with FDA-required labeling, the appropriate US risk information and other required disclosures, properly supported references and the local email template.
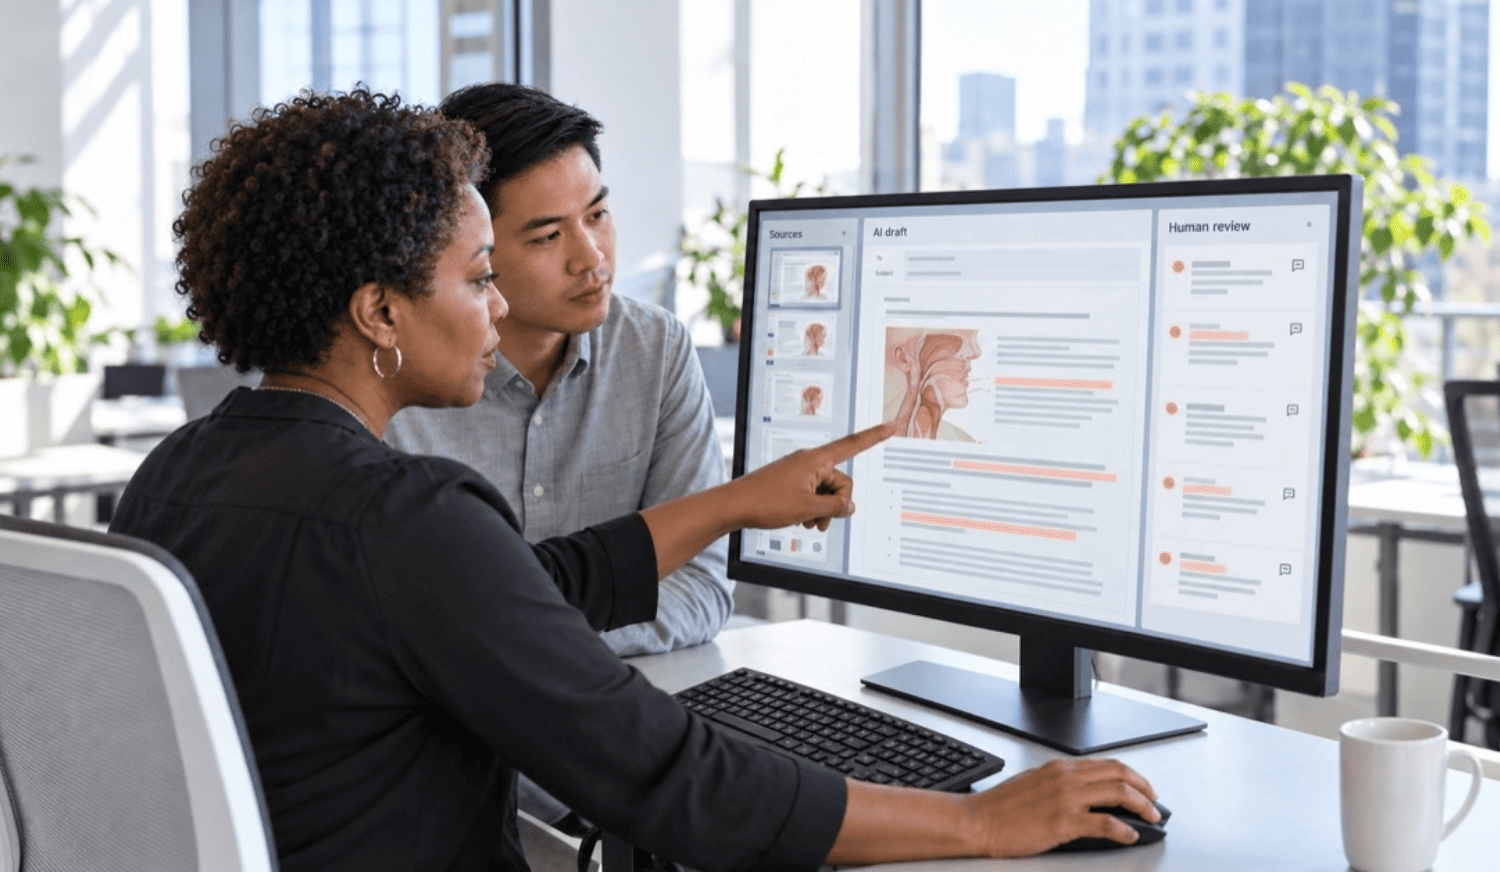
An AI assistant starts with the approved global pack and the company’s US source library. It locks the claims and references that must stay unchanged. It prepares the US draft, inserts the required local elements and marks every change. If a global claim has no direct support in the US sources, the assistant flags it for review rather than rewriting it as fact.
The reviewer receives the draft with a change log and a short list of items that need a decision. If FDA-required labeling changes a week later, the system can identify the affected email and other materials built from the same source. The team no longer needs to search through folders and email threads to find them.
What the software should do
The assistant needs approved global content, current FDA-required labeling, a claims and references library, brand terminology, templates and examples of accepted work. It should separate fixed content from sections open to US adaptation. Every proposed change should point back to a source or rule. Missing or conflicting information should go to a person, never be guessed.
FDA’s Office of Prescription Drug Promotion says prescription drug information should be truthful, balanced and accurately communicated. Its guidance and educational material flag common problems such as unsupported claims, missing or downplayed risks and misleading comparisons. An AI system used in this process should help reviewers find those issues earlier, not make the compliance decision for them.
Human approval stays in the process. The software prepares and documents the draft. Qualified teams make the decisions and release the content.
Start with one recurring asset
Choose one asset family with enough volume to measure, such as HCP emails or approved detail aids. Record preparation time, handoffs, review cycles, common corrections and cost per version before the test begins.
Compare the same measures after the first AI-assisted cycle. How much preparation time changed? Did reviewers receive cleaner drafts? Were fewer versions returned for avoidable corrections? Did the US team publish sooner?
Content localization is one of five connected AI workflows for pharma Commercial and Omnichannel teams. Explore all five use cases.
Sources
Veeva Systems, Building the Future of MLR with AI (2025)
US Food and Drug Administration, Medical Product Communications That Are Consistent With the FDA-Required Labeling – Questions and Answers (2018)
US Food and Drug Administration, The Bad Ad Program
US Food and Drug Administration, Prescription Drug Advertising and Promotional Labeling (Office of Prescription Drug Promotion)